This guide explains Drf Crefito in an evidence-informed, clinical manner—covering what it is used for, how suppliers typically describe it, and how clinicians approach selection. Background information follows in an objective tone, outlining key concepts such as material compatibility, procedural workflow, and quality documentation requirements for safe, effective care.
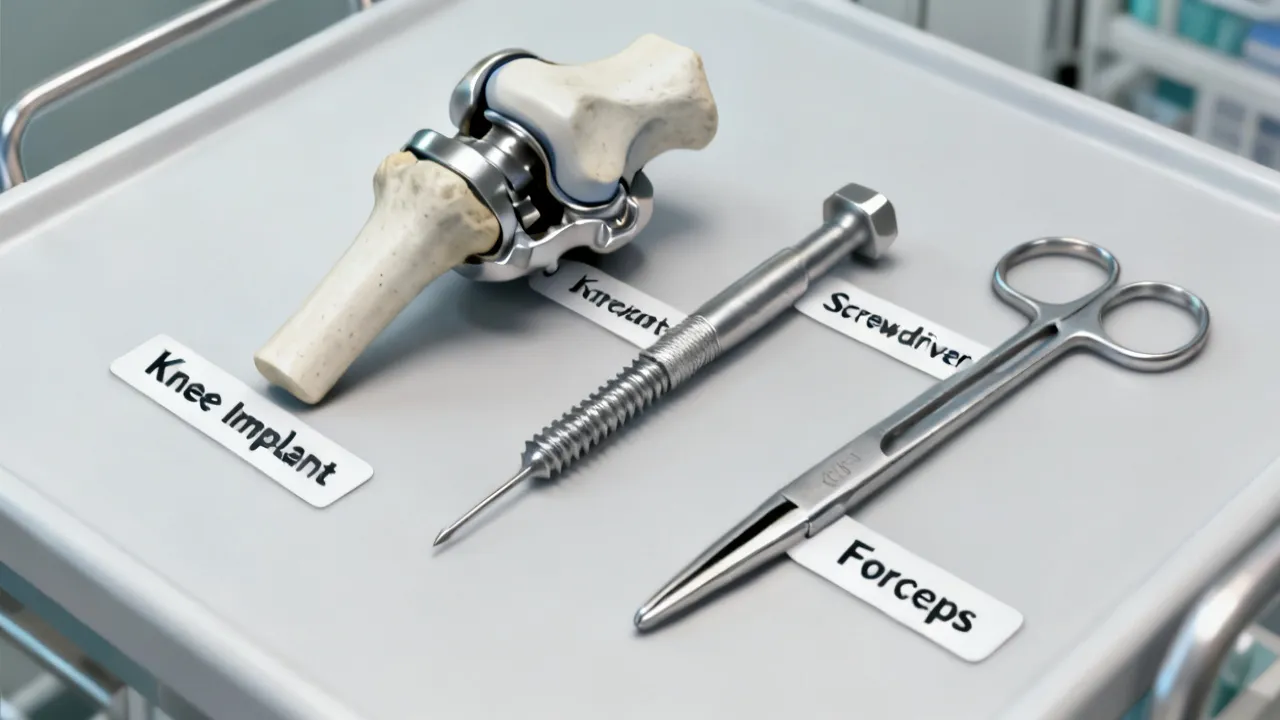
“Drf Crefito” is frequently referenced in orthopedic and musculoskeletal discussions as a brand, model name, or shorthand associated with medical hardware used in treatment planning. In real-world clinical decision-making, however, “what it is called” rarely matters as much as “what it actually is”—the exact product configuration, the verified indications, the materials and design features, the packaging and sterilization status, and the traceability records. Clinicians and procurement teams therefore treat the phrase Drf Crefito as a pointer that must be resolved into a precise identity (manufacturer name, product code, size/variant, and kit contents) and a confirmable evidence set (instructions for use, quality documentation, lot/batch traceability, and labeling).
In practical terms, the safest and most defensible approach is to confirm the exact product configuration and its intended clinical use by reviewing manufacturer documentation and the supporting quality records supplied by the manufacturer or an authorized supplier. This ensures that the device you plan to implant is truly the one you receive, and that it aligns with the surgeon’s procedural plan and the patient-specific anatomical and clinical requirements.
In modern orthopedics, the decision path is typically governed by verifiable information: product labeling, traceability, material specifications, sterilization method, evidence of mechanical performance where relevant, and documented biocompatibility considerations tied to the product’s design and manufacturing controls. When people mention Drf Crefito, the underlying practical question is usually whether the item matches the clinical goal—stabilization, alignment support, reconstructive requirements, or other procedural objectives—while also fitting the patient’s anatomy and fitting within the surgeon’s technique and instrumentation workflow.
From a clinical governance perspective, names can change. Products can be repackaged, bundled into kits, revised in minor ways, or referenced differently across regions and procurement systems. Even if the same general brand shorthand is used, the actual clinical effect of an implant can depend on details like geometry, sizing system, coating or surface finish, and compatibility with specific instruments. That’s why the documentation-first method is not bureaucratic overreach; it’s the mechanism that turns a potentially ambiguous label into a concrete and checkable device identity.
For procurement teams and care coordinators, “supplier details” also matter because supplier roles can vary widely. Some suppliers are primarily distributors with limited customization or limited packaging control; others manage procurement logistics and may provide a documentation packet that must be consistent with institutional traceability and regulatory expectations. Regardless of supplier model, the safest approach is to insist on the same core documents and traceability fields so that the hospital or health system can meet quality management and audit requirements.
From an industry perspective, experienced orthopedic teams evaluate devices in layers rather than by a single identifier. A useful way to structure the assessment for Drf Crefito (or any comparable orthopedic implant reference) is to consider:
These layers translate into practical checks.
1) Clinical fit: Confirm the indications and anatomical suitability. Even when two items share a similar name, differences in design features can be clinically meaningful. A given implant line might have multiple variants: different sizes, different configurations for left/right, different surface coatings, different fixation strategies, or different compatibility boundaries with other components.
2) Quality assurance: Verify traceability and batch/lot information, sterilization status, and packaging integrity. In regulated environments, these details are often validated at multiple points: upon receipt into clinical stock and again as part of operative setup verification. For sterile devices, the barrier integrity and labeling accuracy are particularly important because they affect the ability to maintain sterility through the handling chain.
3) Workflow compatibility: Confirm whether the product is designed to integrate with specific surgical instrumentation and whether the supplier provides implementation guidance aligned with the manufacturer’s instructions. A technically correct implant that cannot be implanted efficiently or correctly with the available instruments can create avoidable delays and procedural variability—an outcome that quality systems generally try to minimize.
In other words, industry teams treat the phrase Drf Crefito as the beginning of an identity resolution process, not the end of it.
When people ask about Drf Crefito “price,” they usually mean the total procurement cost associated with the implant or the related components in the kit required for a procedure. In healthcare procurement, final pricing can depend on many factors beyond the core device itself.
Common cost drivers include the following:
Important: Without a clearly specified region, product code, and authorized supplier context, any published “price” can be misleading or incomparable. Two items that look similar on paper may differ in variant, included components, sterility assurance, or documentation pack. For objective selection, prioritize verified product identity and quality records first; then request a formal quote that itemizes the exact configuration you intend to use.
In many institutions, procurement decision models integrate not only device unit cost but also the “total cost of ownership” from a systems perspective: administrative time for receiving and verifying documentation, the cost of inventory holding, the cost of delayed procedures if availability is uncertain, and the operational time required for compliance checks. As a result, “lowest unit price” is rarely the best indicator of the most efficient and safest procurement outcome.
To reduce risk and ensure traceability, procurement teams commonly request an evidence packet for Drf Crefito (or any comparable orthopedic implant product). The exact requirements can vary by institution and region, but a strong baseline includes:
This documentation-first method aligns with standard healthcare governance: it is not about marketing claims or name recognition, but about repeatability, patient safety, and accountability. When documentation is available and consistent, it supports both clinical confidence and audit readiness.
It’s also useful to clarify the supplier’s responsibility boundaries. For example, whether the supplier can provide documentation for the specific lot being shipped, whether they can confirm sterilization and expiry status at the time of receipt, and whether the supplier can provide an itemized packing list that matches what is described in the purchase order. These details help ensure that the hospital receives exactly what was authorized and planned.
In a typical orthopedic workflow, implant selection is rarely a single isolated decision. Instead, it occurs within a sequence of preoperative evaluation, planning, verification, and postoperative monitoring.
When a product is commonly referenced as Drf Crefito, the practical question for clinicians and coordinators becomes: “Does this exact configuration meet the plan for this patient, and is it delivered with the correct traceability and documentation?”
This phrasing matters because it reflects how quality systems are designed: they aim to reduce the gap between planning and execution. The “name” is just a reference point; the “exact configuration with traceability” is what can be verified and recorded.
Although clinical teams follow local protocols, there are consistent risk management principles across orthopedic practice. These principles can be applied to Drf Crefito in a structured way to reduce avoidable complications related to mismatch, handling errors, or documentation gaps.
This “process discipline” approach is widely endorsed in quality systems because it reduces variation—one of the most common sources of preventable clinical complications. It also supports clear accountability: if an issue arises, teams can trace it to a specific device lot and documentation context rather than relying on memory or informal references.
Additionally, risk management is not limited to the operating room. Procurement and preoperative verification reduce upstream risk. For example, ensuring correct device identity and sterility assurance at receipt helps prevent last-minute substitutions or rushed decisions that can occur if the “wrong configuration” is discovered late.
The following comparison helps teams decide what to prioritize when evaluating Drf Crefito through a procurement lens. This is not a claim about clinical superiority; it is a structured way to align operational checks with governance goals.
| Procurement Priority | What to Check | What Good Documentation Looks Like |
|---|---|---|
| Identity accuracy | Exact product code, size, kit contents, and variant labeling | Clear manufacturer labeling and itemized quote matching the planned implant |
| Traceability | Lot/batch number and device identification for recordkeeping | Lot numbers visible on packaging plus traceability records |
| Sterility assurance | Sterilization method and expiry/shelf-life | Sterile barrier integrity confirmation and expiration dates |
| Regulatory readiness | Market-specific compliance documents | Market-relevant labeling and documentation supplied by authorized channels |
| Clinical workflow alignment | Instrument compatibility and recommended handling instructions | Procedure-aligned instructions consistent with the surgeon’s technique |
| Service reliability | Timely delivery and response capability for urgent replacements | Supplier SLAs, dispatch confirmation, and escalation process |
Notably, each row describes a verification activity that can be audited. When teams can show what they checked, how they checked it, and what evidence they used, it becomes far easier to demonstrate quality control to internal stakeholders and external regulators.
Below is a practical, step-by-step guide that procurement teams and clinical coordinators can use to evaluate Drf Crefito responsibly. While each institution’s policies vary, the logic is broadly applicable: resolve identity → verify documentation → verify packaging → ensure compatibility → record traceability.
As a practical matter, the above process can be integrated into preoperative planning workflows and checklists to reduce cognitive load on individual staff. When the steps are standardized, variation decreases and compliance becomes more consistent.
In many hospitals and orthopedic programs, there are baseline conditions and requirements expected when procurement evaluates implants like Drf Crefito (or any orthopedic hardware). Typical expectations include:
Many of these expectations can be supported with evidence: photos of packaging labels during receipt, scanned documentation packets, and device history record entries tied to specific lots. Over time, this creates a reliable audit trail.
Orthopedic implant selection is often discussed using brand names and shorthand references because they are easy to communicate in everyday clinical and procurement conversations. Yet clinical decisions depend on standardized information: mechanical design features, materials, manufacturing quality systems, and repeatable performance characteristics. For that reason, healthcare organizations implement procurement safeguards designed to confirm identity, traceability, and sterile integrity.
When buyers discuss Drf Crefito, the practical concerns typically include how the product is described in catalog materials, how suppliers represent specifications, and whether the final delivered item matches what was requested. This is why objective selection emphasizes verified documentation rather than informal descriptions.
Regarding “price,” procurement leaders generally treat cost as one variable within a larger decision model. Total cost considerations can include inventory holding, delivery reliability, compatibility-related workflow costs, and the administrative effort required to maintain traceability and compliance. A lower unit price may be offset by higher operational costs if the device requires extra coordination, experiences delivery variability, or creates documentation verification delays.
In addition, many hospitals are aware of how naming ambiguity can create downstream risk. When an implant is referenced vaguely, the chance of ordering a variant that is “close enough” but not identical increases. Even small differences can impact instrument needs, surgical steps, and the completeness of the kit delivered. Governance systems are designed to prevent those near-miss events.
Healthcare device procurement is shaped by quality management principles and regulatory expectations. Many organizations rely on established quality systems and risk management methods for medical devices. These methods focus on:
In day-to-day operations, these governance goals translate into concrete behaviors: verifying product identity at receipt, ensuring sterile packaging integrity, recording lot/batch information accurately, and maintaining documentation packets that can be audited. The standards do not usually direct a specific brand choice, but they define how the quality system should function so that any approved brand or product can be used responsibly.
For authoritative guidance, organizations often reference materials from recognized regulatory bodies and standard-setting organizations. Two widely used sources in medical device governance include:
Sources for further reading: ISO standards materials (ISO 13485) and FDA medical device information pages and guidance library. These are governance-level references and do not validate any specific brand; they outline how quality systems and documentation expectations are structured.
When institutions align their procurement checklists with such standards, they reduce variation and strengthen accountability. This is particularly relevant when a product reference like Drf Crefito can be misunderstood across teams. The standard-driven workflow helps make the decision reproducible and auditable.
Drf Crefito is commonly used as a reference name for a medical orthopedic device system or component. Because naming can vary by market, product line, or kit configuration, the exact item should be confirmed using manufacturer identity and product code details. In other words, the phrase is usually a label that must be mapped to a specific manufacturer product and variant.
Ask the supplier for the exact manufacturer product code, size options, kit contents, and applicable instructions for use for the configuration you intend to purchase. Then verify that the quote and the received packaging match those details, including lot/batch identifiers and sterility labeling. A strong confirmation process includes receiving/inspection checks and documentation verification—not only a “we quoted the right thing” assumption.
Yes. Pricing can change based on configuration, packaging type, distribution channel, delivery timelines, and whether the supplier provides additional services or administrative support. For reliable comparison, request itemized quotes that specify the exact product configuration, kit contents, and documentation set. Comparing “brand-level” prices without configuration-level alignment often leads to inaccurate conclusions.
Typically, you should request manufacturer identity, traceability/lot details, sterility assurance information, relevant labeling, and instructions for use for the specific configuration. The exact document set depends on your local regulatory environment and institutional policy. In many settings, the “minimum viable” documentation includes instructions for use and lot traceability sufficient for patient-level device history records.
Yes. Clinical requirements depend on the procedure and your surgeon’s protocol. Generally, teams must ensure correct sterile handling, instrument compatibility, and accurate device documentation in patient records. Follow the manufacturer instructions and institutional surgical checklists, and ensure staff are trained on device-specific steps when required.
Use a documentation-first vetting process: confirm identity and configuration, verify traceability and sterility labeling upon receipt, ensure authorized sourcing where possible, and maintain consistent recordkeeping for each device used. Additional risk reduction comes from standardizing checks (so they are not dependent on individual staff memory) and quarantining items that have unclear labels or documentation.
You can compare based on objective criteria—such as configuration, verified specifications, packaging/sterility status, traceability provisions, and instrument compatibility—rather than brand names alone. Any comparison should be anchored to manufacturer documentation and surgeon-specific requirements. A fair comparison typically compares equivalent configurations and kits, not just similar indications.
Ultimately, the responsible way to handle Drf Crefito—whether you are a procurement coordinator, clinical administrator, biomedical engineer, or clinician—is to transform “brand reference” into measurable criteria. By focusing on identity accuracy, traceability, sterile assurance, documentation completeness, and workflow compatibility, teams reduce ambiguity and support safer, more consistent outcomes.
If you share the exact product code or the configuration you’re evaluating (and the authorized supplier context), it becomes possible to build a more tailored procurement checklist and a decision matrix aligned with your intended orthopedic application, including the specific documents to request, the compatibility checks to perform, and the receipt/inspection criteria that should be logged before the device is used.
Striking the Perfect Balance: Navigating Premiums and Out-of-Pocket Expenses in Senior Insurance Plans
Explore the Tranquil Bliss of Idyllic Rural Retreats
How to Make Lasting Memories at Disneyland Attractions
Affordable Phones and Plans for Seniors
Affordable Full Mouth Dental Implants Near You
Unlock the Top Kept Secrets to Finding Your Ideal Dentist for Flawless Dental Implant Results!
Discovering Springdale Estates
Unveiling RS Sul Telecom Services
The Guide to Car Trading
Striking the Perfect Balance: Navigating Premiums and Out-of-Pocket Expenses in Senior Insurance Plans
Finding an optimal mix of insurance premiums and direct medical costs is crucial for effective healthcare budgeting. This discussion will delve into tactics to achieve a balance that aligns with your finances.
Unlock the Top Kept Secrets to Finding Your Ideal Dentist for Flawless Dental Implant Results!
Selecting an adept dentist is critical for ensuring the efficacy and your satisfaction during a dental implant procedure. This in-depth guide is designed to assist you in finding a dental expert who meets your personal requirements, guaranteeing that your path to dental implant success is seamless and fruitful.
Protect Your Perfect Smile: Master the Art of Defending Your Teeth Against Acid Attacks
Though they may tantalize the palate and invigorate the senses, foods and drinks with high acid content are a double-edged sword with regard to oral wellness. Continual exposure to these substances progressively strips away the protective enamel from our teeth, which can lead to increased sensitivity, changes in coloration, and a variety of other dental complications. This composition offers insight into relishing these zesty favorites while safeguarding your teeth from damage.
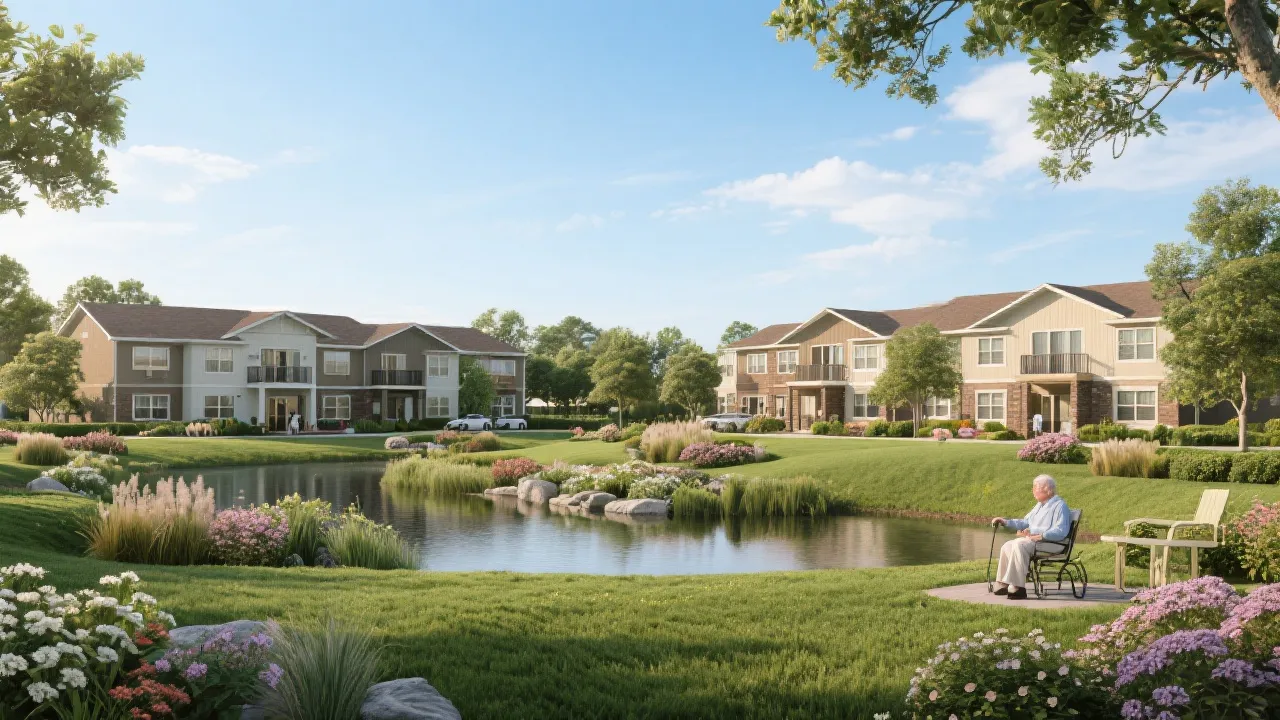
Discovering Heartis Eagle Mountain
Heartis Eagle Mountain is a prominent senior living community that offers a range of services and amenities aimed at enhancing the quality of life for its residents. Situated in a peaceful setting, it combines exceptional care with engaging activities, providing a holistic environment that caters to seniors' physical, emotional, and social needs.
Unveiling the Top Dental Implant Options for Seniors: Transform Your Smile with the Ultimate Guide to Restored Radiance
The significance of oral health cannot be understated for the elderly population as they are commonly afflicted with tooth loss due to a multitude of factors, including decay and gum disease. Dental implants have risen to prominence as a prime method of teeth replacement, acclaimed for their long-lasting nature, functional prowess, and striking similarity to our own teeth. This discussion delves deeply into the realm of dental implants to demonstrate why they are considered the optimal choice for aged individuals aiming to enhance their smiles and overall life quality.
Boost Your Defenses: 20 Proven Strategies for a Stronger Immune System
Having a robust immune system is essential to protect our bodies from harmful microorganisms and preserve our general health. Although some elements that affect our immunity are out of our hands, many lifestyle decisions can help us enhance our immune defenses naturally. This piece delves into 20 practical methods to empower your immune system, increasing your capacity to ward off diseases and sicknesses.